Put tert-butyl bromide, (CH3)3C–Br, in warm water and its C–Br bond breaks on its own. The bromine leaves with both electrons of the bond, as bromide ion. The carbon it leaves behind keeps only three bonds and carries a positive charge. That species, (CH3)3C+, is the tert-butyl cation.
A carbocation is any carbon with three bonds, no lone pair and a positive charge. Carbocations are short-lived, but many of the reactions ahead pass through one. Most questions about them come down to two skills: judging how stable a particular cation is, and spotting when it will rearrange into a more stable cation before a nucleophile can reach it.
What it is, and how to check your drawing
Count the electrons on the charged carbon of the tert-butyl cation. Carbon brings four valence electrons. In the cation it owns no lone-pair electrons and one electron from each of its three bonds, so the formal-charge sum from Foundations gives 4 − (0 + 3) = +1.
Those three bonds hold six electrons, two short of an octet. That shortfall is what makes a carbocation so reactive: anything with a pair of electrons to offer is drawn to it.
Its shape: flat, with an empty p orbital
A carbon with three groups and no lone pair has only three electron domains, and they spread as far apart as they can. So the cation is trigonal planar, with bond angles near 120°, and its carbon is sp2-hybridized, as the Hybridization section of Foundations showed for CH3+. The three sp2 orbitals make the three bonds. The fourth orbital, an unhybridized p orbital, is empty. It stands at right angles to the plane of the bonds, with one lobe above the plane and one below it.
The carbon in tert-butyl bromide was tetrahedral and sp3, so forming the cation flattens it. A flat cation has two identical faces. Nothing blocks either lobe of the empty orbital, so a nucleophile, a species that donates an electron pair to form a bond, can reach the carbon from above or from below with equal ease.
That matters when the carbon was a stereocenter. Take the butan-2-yl cation, CH3CH+CH2CH3, which 2-bromobutane gives when its bromide leaves. Bromide returning from above gives one enantiomer of 2-bromobutane; bromide returning from below gives its mirror image. The two form in equal amounts, a 50:50 mixture that Stereochemistry called a racemic mixture. The handedness of the starting material is lost the moment the flat cation forms, not scrambled at some later step.
Where they come from
Two routes make nearly all carbocations, and both appear in the figure below.
A leaving group departs with its bonding pair. This is the event the Leaving groups section was about. In tert-butyl bromide the C–Br bond breaks heterolytically, meaning one atom takes both of its electrons. Bromine leaves as bromide ion, and the carbon keeps three bonds and the charge. A leaving group that starts out positive, such as –OH2+, leaves as a neutral molecule instead. How fast this step goes depends on two things together: how stable the leaving group is once it has gone, and how stable the cation is once it has formed.
A π bond takes a proton. The two π electrons of a C=C are loosely held and exposed above and below the bond, so they can attack a proton, such as the H of H3O+. In 2-methylpropene, (CH3)2C=CH2, the H adds to the CH2 end. The other alkene carbon gets no share of the π pair and becomes the cation, which is the same tert-butyl cation that route 1 gives. Adding the H to the other end would leave a primary cation, and the next section explains why that is much worse. The reactions themselves belong to Alkenes & Alkynes.
Stability: alkyl groups help
Compare four cations: CH3+, CH3CH2+, (CH3)2CH+ and (CH3)3C+. Their charged carbons carry zero, one, two and three carbon groups, and they become more stable in that order. The number of carbons bonded to the charged carbon is its degree: one makes it primary (1°), two secondary (2°), three tertiary (3°). The methyl cation, with none, gets no degree and is simply called methyl. So the order is 3° > 2° > 1° > methyl.
The degree counts only the carbons attached to the charged carbon itself, not every carbon in the molecule. The pentyl cation, CH3CH2CH2CH2CH2+, has five carbons but is still primary.
Two effects produce the order, and both push the same way. Hyperconjugation is the larger effect. In the ethyl cation, CH3CH2+, the CH3 group next door has three C–H bonds. When one of them lines up parallel to the empty p orbital, its electron pair spreads a little into that orbital. No bond breaks and no new bond forms; the pair simply spends part of its time nearer the positive carbon. That moves some of the positive charge off the cationic carbon and onto the neighboring C–H bond. The CH3 group rotates freely, so each of its C–H bonds takes a turn.
More alkyl groups mean more C–H bonds next door: none for the methyl cation, three for ethyl, six for isopropyl and nine for tert-butyl. C–C bonds on a neighboring carbon donate in the same way, so a neighbor with no hydrogens still helps.
Induction, the push or pull of electrons through σ bonds (see the Electron-rich vs. electron-poor atoms section), is the smaller effect. An alkyl group has a bigger, looser electron cloud than a hydrogen atom, which makes it more polarizable. So it pushes electron density along the σ bond toward a neighboring positive charge more readily than an H does. Neither effect moves the charge fully onto another atom; both only spread it out.
The effect on rates is large. In solvolysis, a reaction in which the solvent itself is the nucleophile, a tertiary alkyl bromide warmed in aqueous ethanol ionizes to a carbocation. A tertiary bromide reacts a million or more times faster than methyl bromide, the exact figure depending on the solvent. Read that number with care. Methyl and primary bromides hardly ionize at all, and the slow rate they do show comes mostly from a different mechanism. What the ratio really says is that tertiary bromides ionize and methyl bromide does not.
Resonance beats substitution
Now compare the allyl cation, CH2=CH–CH2+, with the ethyl cation. Both are primary, yet the allyl cation is far more stable. Its empty p orbital sits right beside the π bond, so the π electrons can move toward it, and the charge ends up shared between the two end carbons. The Resonance section of Drawing Molecules & Moving Electrons drew this with curved arrows.
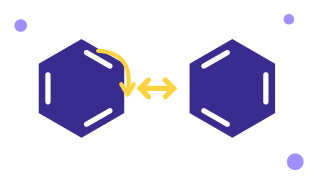
A cation next to a C=C is called allylic, and one next to a benzene ring is called benzylic. In a benzylic cation the charge spreads onto three ring carbons as well as the outside one: the two ring carbons beside the attachment point and the one across the ring from it. Hyperconjugation only lends a little of the charge to neighboring bonds, while resonance moves it onto other atoms outright. That gives a ranking worth memorizing: a primary benzylic cation is about as stable as a tertiary alkyl one, and a primary allylic cation is about as stable as a secondary alkyl one. So count the groups on the charged carbon first, then check for a neighboring π bond or ring, which moves the cation one or two places up the order (the ladder figure below sums it up).
A lone pair next door does better still. In CH3O–CH2+, a lone pair on oxygen can move in to form a C=O π bond. That gives a second resonance structure in which every atom has an octet and the positive charge sits on oxygen. Oxygen is electronegative and carries the charge reluctantly, but a structure with full octets everywhere beats one with a six-electron carbon, so this cation is more stable than even a tertiary alkyl cation. A cation with oxygen on the charged carbon is an oxocarbenium ion, the intermediate in acetal formation in Carbonyl Chemistry. The nitrogen version, with a C=N+ π bond, is an iminium ion. It is more stable again, because nitrogen is less electronegative than oxygen and shares its lone pair more readily.
Vinyl and aryl cations: the empty orbital points the wrong way
Two kinds of cation sit on or beside a π system and still get no help from it. A vinyl cation has its charge on a carbon of a C=C, as in CH2=C+–CH3. That carbon has only two groups, so it is linear and sp-hybridized. An sp orbital is half s character and holds its electrons close to the nucleus, which makes an sp carbon a poor place for a positive charge.
The carbon’s own π bond cannot rescue it. The vinyl carbon uses one of its two p orbitals for the π bond, and the empty one is the other p orbital, at right angles to the first. So the empty orbital is perpendicular to the π bond, the two cannot overlap, and the π electrons cannot spread into it.
An aryl cation has its charge on a carbon of a benzene ring. The ring holds that carbon at sp2 and will not let it straighten out, so its empty orbital is one of the sp2 hybrids. That hybrid lies in the plane of the ring and points outward, while the ring's π cloud sits above and below the plane. Again the empty orbital and the π electrons are at right angles, and they never meet.
So an ordinary vinyl or aryl halide does not ionize, and no mechanism that needs one of these cations can start from it. This is also why substitution at a benzene ring carbon has to follow other mechanisms, and Aromatic Chemistry covers them. An aryl cation can still form when the leaving group is exceptionally good: an aryl diazonium salt, which you will meet in Aromatic Follow-Through, loses N2 gas on warming and leaves one behind.
Rank these from most to least stable: (i) the tert-butyl cation, (CH3)3C+; (ii) the benzyl cation, C6H5CH2+; (iii) CH3O–CH2+; (iv) the phenyl cation, C6H5+.
Sort them by what stabilizes the charge, not by size. In (iii) an oxygen lone pair sits on the charged carbon, so the cation has a resonance structure with a C=O+ π bond and an octet on every atom. That is the strongest stabilization on the list, even though no carbon is attached to the charged carbon. (ii) is only primary, but the ring spreads its charge over four carbons, which puts it level with a tertiary alkyl cation. (i) has nine C–H bonds next door, the best an alkyl cation can do. (iv) is an aryl cation: its empty sp2 orbital lies in the ring plane and gets nothing from the π system.
Answer: (iii) > (ii) ≈ (i) >> (iv). There are two traps. (iv) contains a benzene ring and looks stabilized, but the ring cannot help it. (iii) has the fewest carbons of the four and still comes out on top.
Rearrangements: cations do not stay put
Treat 3,3-dimethylbutan-2-ol, (CH3)3C–CH(OH)–CH3, with HBr. Swapping the OH for Br in place would give 3-bromo-2,2-dimethylbutane. The main product is 2-bromo-2,3-dimethylbutane instead, because a methyl group has moved to the neighboring carbon. The cation rearranged before bromide reached it.
Number the chain from the end nearer the OH. The OH is on C2. C3 is bonded to C2, C4 and two methyl groups, so it has no hydrogen.
Step 1 — activate the OH. Hydroxide is a poor leaving group, so the acid first puts a proton on the oxygen. The group that leaves is then a neutral water molecule, just as in the Leaving groups section.
Step 2 — ionize. Water leaves and takes the C–O bonding pair with it. C2 keeps three bonds, to C1, to C3 and to its H, so the cation is secondary.
Step 3 — look at the neighbors. C1 is a methyl group. Moving one of its hydrogens to C2 would put the charge on C1, a primary carbon, which is uphill, so nothing happens there. C3 has no hydrogen, but it does have methyl groups. Moving one of them to C2 leaves the charge on C3, which still holds a methyl group, C4 and C2. That is a tertiary cation, so this shift happens.
Step 4 — the shift. The arrow's tail sits on the C3–CH3 bond and its head on C2. The methyl group moves to C2 with its two bonding electrons, and the charge ends up on C3.
Step 5 — capture. Bromide bonds to C3. Numbered afresh from the end nearer the Br, that carbon is C2, so the product is 2-bromo-2,3-dimethylbutane.
This substrate is a favorite exam trap. The carbon next to the cation has no hydrogen, so a student who looks only for a hydrogen to move finds nothing and gives the unrearranged product.
The move in step 4 is a 1,2-shift. A group on the carbon next to the positive carbon moves across to it, taking its bonding pair along, and the positive charge ends up on the carbon the group just left. A carbocation lives only briefly, but usually long enough for this to happen before a nucleophile arrives.
Two groups move this way routinely. A 1,2-hydride shift moves an H together with its bonding pair; the H travels as hydride, H−, because it takes both electrons. A 1,2-methyl shift moves a CH3 in the same way, and other alkyl groups can move too. In the 3-methylbutan-2-yl cation below, (CH3)2CH–CH+–CH3, the neighboring carbon has an H, and moving it turns a secondary cation into a tertiary one. The neopentyl cation, (CH3)3C–CH2+, has no H on the carbon next door, so a methyl group moves instead.
The arrow is the part most often drawn wrong. Its tail goes on the bond that moves, the C–H or C–CH3 bond, rather than on the H or CH3 itself or on the positive carbon, because what moves is a bond and its two electrons. Its head goes on the positive carbon.
The rule for predicting a shift is short. A cation shifts when one shift gives a more stable cation. It never shifts to a less stable one, and once it reaches the most stable cation on offer, it stops. What decides the question is the cation the shift leaves behind, not whether the moving group is H or CH3. Only a group on the immediate neighbor can move; a hydrogen two carbons away is out of reach.
Exams grade by that rule, and it is right for shifts to a better or a worse cation. A cation can also shift to an equally stable one, for example from one secondary carbon to another, and the reaction then gives a mixture; exam answers usually leave such shifts out.
(Chloromethyl)cyclobutane is a cyclobutane ring carrying a CH2Cl group. Warmed in water, it gives cyclopentanol: a five-membered ring from a four-membered starting material.
Step 1 — chloride leaves. On paper this leaves a primary cation on the outside CH2. A primary cation is very poor, which is why the reaction is slow.
Step 2 — two shifts are possible. The ring carbon next to the CH2+ carries one hydrogen. A hydride shift would give a tertiary cation, 1-methylcyclobutyl, still sitting on a strained four-membered ring. That ring carbon is also bonded to two other ring carbons. If one of those C–C bonds shifts instead, the carbon at its far end (labeled a in the figure) moves off the ring carbon and bonds to the CH2. Two things improve at once. The charge lands on a ring carbon, which makes it secondary, and the ring grows from four members to five. This is a 1,2-alkyl shift in which the moving group is part of the ring. A four-membered ring carries a lot of angle strain (Alkanes & Conformations) and a five-membered ring very little, so the ring-expansion shift wins.
Step 3 — capture. The result is the cyclopentyl cation. Water bonds to it, a second water molecule removes the proton from the new O–H, and the product is cyclopentanol.
Drawing this as “ionize, then shift” is useful bookkeeping but not quite what happens. A free primary cation is too unstable to last, so the ring bond starts to move while the chloride is still leaving, in one motion. That is why this primary substrate reacts at all. Whenever a rearrangement would also relieve ring strain, look for it before you look for a hydride shift.
Why a more stable cation also forms faster
Stability describes a species, while rate depends on a transition state. The Hammond postulate from the previous section links the two. Forming a carbocation is strongly uphill, because a bond breaks and, from a neutral starting material, two charged pieces appear. So the transition state for that step comes late and looks much like the cation itself, and anything that lowers the cation's energy lowers the barrier to it by nearly as much.
That is why carbocation stability predicts more than which intermediate lasts longest. It also predicts which reaction goes faster, which of two possible cations forms, and therefore which product you isolate. Many arguments later in the course rest on this link: which way an acid adds across a C=C, which substrates react by the ionizing substitution and elimination mechanisms, and which charged fragments dominate in mass spectrometry, the method that sorts the charged pieces of a broken molecule by mass.
What carries forward
The next section, radical halogenation, ranks its intermediates by the same hyperconjugation argument, with one electron in the p orbital instead of none. The ionizing substitution and elimination mechanisms in Substitution & Elimination run through a carbocation, and rearrangements explain why their products sometimes have a different carbon skeleton from the starting material.